August 25, 2026
Written by Austin Tyner
Clinical Review by Valerie Schwartz
When a deficiency letter arrives from the U.S. Food and Drug Administration (FDA) during substantive review of a Premarket Tobacco Product Application (PMTA), .the application is placed on hold pending a response within the window stated in the letter, commonly 90 days. The applicant may, during this time, attempt to clarify the information presented in the submission or to contextualize submitted product data. This will, in most cases, prove insufficient because deficiencies typically reflect gaps in the underlying evidence rather than in how clearly that evidence was presented. Missing studies, inadequate product characterization, or development and quality processes that fail to generate necessary data cannot be corrected through narrative alone. Compounding the problem, the length of the response window may not allow enough time to conduct additional studies. Deficiency letters serve a specific function in the FDA PMTA review process. Knowing what they contain, why applicants receive them, and how to respond puts an applicant in a stronger position to support an Appropriate for the Protection of the Public Health (APPH) determination.
Understanding Deficiency Letters in the FDA Review Sequence
The FDA reviews a PMTA in several successive stages, and an application can be found insufficient at any of them. Each stage carries its own FDA action and requires applicant response. These stages are, in order, acceptance review, filing review, substantive application review, and finally a marketing order decision.
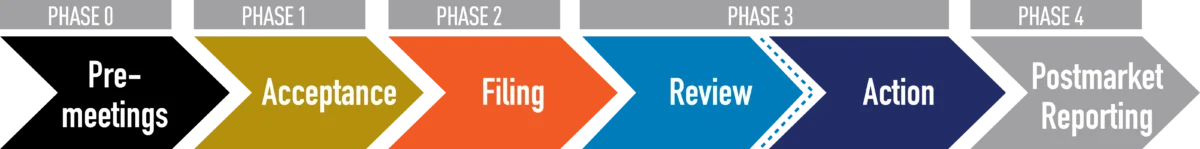
PMTA review process, courtesy of fda.gov
A deficiency letter results from gaps or inaccuracies identified during substantive review, when FDA subject matter experts examine the application in full to determine whether all necessary information is present. A deficiency letter is distinct from other correspondence stemming from issues elsewhere in the review process, such as a refuse to accept letter or a refuse to file letter.
The differences between a deficiency letter and other review-related letters are stated as follows:
| Letter Type | Review Process Step | Criteria for Refusal |
|---|---|---|
| Refuse to Accept (RTA) | Acceptance Review | Product does not fall under CTP jurisdiction, or the statutory and regulatory requirements of the application are not met, based on Section 910 of the FD&C Act and the criteria set forth in 21 CFR 1114.27(a)(1). |
| Refuse to File (RTF) | Filing Review | Application does not contain sufficient information to permit a substantive review in accordance with 21 CFR Parts 1100, 1107, and 1114. |
| Deficiency Letter | Substantive Application Review | Additional information is needed to complete the scientific review. Specifics are listed in the deficiency letter. |
Common ENDS Product Deficiencies Identified by Accorto
In substantive application review, many types of informational gaps may be identified by the FDA and detailed in a deficiency letter. The FDA holds every applicant product to the APPH standard, meaning the applicant product's public health benefit must be greater than its risk to the population as a whole, including users and nonusers. The FDA will not issue a marketing granted order if that standard is not met, and the FDA can only determine whether the applicant product meets the APPH standard when it is given an application that addresses each question asked with organized, product-specific data drawn from appropriate rigorous testing.
According to subject matter experts at Accorto, the most commonly observed recurring deficiencies are gaps in the behavioral science sections of the application, specifically Tobacco Product Perception and Intention (TPPI) studies and actual-use studies. Applicants frequently substitute general category research or consumer marketing data for product-specific evidence. Importantly, the FDA does not say that every PMTA automatically requires a TPPI study or an actual-use study of a particular design. What testing is needed depends on the specific questions the FDA must answer to reach an APPH determination for that product. TPPI studies evaluate consumer product perceptions and how those perceptions may influence use behaviors. Actual-use studies examine how consumers use the product under real-world or near-real-world conditions. Each answers a different question relevant to the APPH evaluation. Category-level research and marketing data may provide useful context, but they rarely answer the product-specific questions the FDA is asking — which is how they become PMTA deficiencies.
A second gap Accorto observes frequently is analytical chemistry that characterizes the e-liquid without characterizing the aerosol. Analytical chemistry is foundational to a PMTA because it allows reviewers to assess the constituents and potential toxicants users may be exposed to, a critical component of evaluating APPH. In ENDS devices, the process of heating and aerosolizing an e-liquid can generate new compounds or change constituent concentrations. As a result, e-liquid testing alone cannot adequately characterize inhalation exposure. Aerosol testing provides product-specific evidence of e-liquid transfer efficiency to the aerosol and of other byproduct constituents to which a user may be exposed. The FDA expects applicants to characterize aerosol constituents, and its published list of Harmful and Potentially Harmful Constituents (HPHCs) in tobacco products is a useful starting point for identifying analytes for targeted aerosol testing.
A third gap Accorto sees regularly is toxicological evidentiary support for inhalation of the product ingredient set. This deficiency is an example of product-specific testing being set aside in favor of more generalized information. Applicants, in this instance, may provide constituents or general safety information without adequately specifying toxicology-specific information such as inhalation-specific toxicity, expected exposure levels, potential interactions among ingredients, or risks associated with thermal degradation products. Because safety conclusions from oral, dermal, or food-use data do not necessarily translate to inhalation exposure, it is important to provide evidence that is relevant to the product's actual route and conditions of use.
Another critical component of a PMTA where Accorto has observed deficiency gaps is in the manufacturing and quality documentation. The APPH standard requires a PMTA to demonstrate that the applicant product can be manufactured consistently and in a controlled manner. Gaps in documentation covering specifications, batch controls, ingredient qualification, process validation, stability, or the quality management system can create uncertainty as to whether the product evaluated in the application is representative of the product that will ultimately reach consumers.
General Contents of a Deficiency Letter
Deficiency letters are delivered via email and generally include at least one major issue and place the marketing application on hold pending the FDA's receipt of the requested additional information. Minor deficiencies may still be included in deficiency letters when related to the resolution of substantive issues.
A deficiency letter may, in general, include:
- What was submitted: Acknowledgment of the information submitted by the applicant, including references to sections, page numbers, or tables where appropriate.
- Identification of a specific issue or concern with information that was submitted, is missing, or is inadequate.
- Statement of basis for the deficiency that includes the effect or impact of the specific issue or concern on the marketing authorization decision, and, when available, applicable, and relevant, a specific reference.
- Explicit request for the additional information needed to address the issue and potential alternate ways of satisfying the issue, if applicable.
The Accorto team notes that the deficiency letter will state the window within which a response is necessary.
The practical cost of a deficiency can extend beyond the response window. Under 21 CFR 1114.9(b)(1), the FDA will restart the 180-day review period if a deficiency response constitutes a major amendment, such as an amendment containing significant new data from a previously unreported study, detailed new analyses of previously submitted data, or substantial new manufacturing information. Therefore, addressing a deficiency that requires further testing to fill evidentiary gaps may consume some or all of the response window. If the submission of resulting data is a major amendment, then that 180-day review period will be restarted by the FDA upon submission receipt.
Accorto’s Approach To Deficiency Letter Response
The following format may be appropriate for applicants responding to deficiencies:
- Restate the identified Agency issue; and
- Provide one of the following:
- the information or data requested;
- an explanation why the issue does not affect or impact the marketing authorization decision; or
- alternative information and an explanation describing why the information adequately addresses the issue.
Applicants should provide the deficiency number and an identical restatement of the Agency's question when responding to a particular deficiency. If you are responding to a follow-up question from a previous deficiency, include both the original deficiency and the follow-up question. Organize the information with a table of contents, a list of figures, and a list of tables, as necessary to facilitate ease of review. In response to deficiencies, include a description or justification of how the information adequately addresses the Agency's concerns. When providing a declaration of conformity to FDA-recognized consensus standards in lieu of data, the applicant should identify the standard, its revision date, applicable sections, and any deviations from the standard.
Note on submission of response: after November 14, 2026, eSubmitter will no longer be available for PMTA and SE submissions. Applicants should be prepared to use the web-based forms in CTP Portal NextGen before that date.
How Accorto Can Help
At Accorto, our team can help PMTA applicants identify and address evidentiary gaps before they become deficiencies. For applicants with existing deficiency letters, our team can evaluate the FDA’s requests and organize a response that directly answers each concern, giving applicants’ products their best chance at an APPH determination.
Sources
Premarket Tobacco Product Applications. 21 CFR Part 1114.
https://www.ecfr.gov/current/title-21/part-1114
Premarket Tobacco Product Applications and Recordkeeping Requirements. 86 FR 55300.
https://www.federalregister.gov/documents/2021/10/05/2021-21011/premarket-tobacco-product-applications-and-recordkeeping-requirements
U.S.Food and Drug Administration Harmful and Potentially Harmful Constituents (HPHCs)
https://www.fda.gov/tobacco-products/products-ingredients-components/harmful-and-potentially-harmful-constituents-hphcs
U.S. Food and Drug Administration. (n.d.). Premarket Tobacco Product Applications.
https://www.fda.gov/tobacco-products/market-and-distribute-tobacco-product/premarket-tobacco-product-applications
U.S. Food and Drug Administration. (n.d.). Preparing and Submitting a Premarket Tobacco Product Application.
https://www.fda.gov/tobacco-products/premarket-tobacco-product-applications/preparing-and-submitting-premarket-tobacco-product-application
U.S. Food and Drug Administration. (n.d.). Submit and Reference Tobacco Product Master Files
https://www.fda.gov/tobacco-products/market-and-distribute-tobacco-product/submit-and-reference-tobacco-product-master-files
Austin Tyner
Document Specialist
Austin Tyner is Accorto’s document specialist, supporting the quality and publishing team with document onboarding, formatting, and FDA submission readiness. She holds a BA in English and Comparative Literature, with a minor in Public Policy, from the University of North Carolina at Chapel Hill.

Valerie Schwartz
Regulatory Specialist I
Valerie Schwartz, our regulatory affairs manager, began her career as a scientist at the United States Geological Survey, specializing in reconstructing historical climate records. Transitioning into industry, Valerie excelled at Juul Labs, where she contributed as an R&D analytical chemistry scientist, driving new product development initiatives. She later advanced into Regulatory Operations, spearheading non-clinical regulatory science projects. Valerie holds a Master of Science degree in Geosciences from San Francisco State University and earned her Bachelor’s degree in Geosciences and Chemistry from Skidmore College.